Getting started with GreenFold
Help & worked examples
Inspect precomputed MSAs, prepare model inputs, and understand the evidence returned for each protein chain.
Supported models: AlphaFold 2 · AlphaFold 3 · Boltz · OpenFold3 · Protenix
What GreenFold returns
GreenFold reuses indexed MSAs for reviewed human proteins and matches submitted protein sequences to eligible reference entries. It prepares input packages for AlphaFold 2, AlphaFold 3, Boltz, OpenFold3 and Protenix.
The server returns alignments, reference-match information and model input packages. Run structure prediction separately in your chosen model environment. Any AlphaFold DB structure displayed in the interface is a reference structure from AlphaFold DB.
Public access requires no registration or API key. Reference matching and GreenFold MSA reuse depend on the indexed human reference set. A sequence outside that set may need native processing or the supported ColabFold partial workflow.
Quick start: prepare a model input
Choose your model below. All five examples use the same 44-residue reference protein, P63313, in the model's native format. AlphaFold 3 is the default selection in Prepare.
| Model | Native input | Worked example | Guide |
|---|---|---|---|
| AlphaFold 2 | FASTA | Load input · Open output | AF2 guide |
| AlphaFold 3 | JSON | Load input · Open output | AF3 guide |
| Boltz | YAML | Load input · Open output | Boltz guide |
| OpenFold3 | JSON | Load input · Open output | OpenFold3 guide |
| Protenix | JSON | Load input · Open output | Protenix guide |
- Select Load input for your model in the table above.
- Inspect the original FASTA, JSON or YAML preview. To edit it, change the protein entity; the preview then follows the entity editor. You can also import your own file with Start from an existing model input.
- Select Prepare model input ZIP. GreenFold matches protein chains, stages eligible MSAs and rewrites the native input.
- Read the chain table. Check the UniProt assignment, identity, MSA mode, pairing and warnings before proceeding.
- Download and unpack the ZIP. Follow
RUN_COMMANDS.mdin the appropriate model environment. The ZIP contains prepared inputs, not a newly predicted structure.
Select Open output to inspect a saved result immediately. The input preparation guide explains conversion, pairing and fallback for each model.
To try the examples already built into the editor, open Start from an existing model input, choose Load an example for this model, then Read into entities. These small examples demonstrate file formats; protein/ligand combinations in them do not establish binding.
Worked examples with interactive output
Each output below was generated by the public service from the downloadable input. Open it in the same Prepare interface used for your own results, inspect the report and commands, or download the ZIP. Loading an input lets you edit it and submit a fresh request.
The reference example below is available for all five models; choose one to inspect its native input, prepared output and run commands. The additional walkthroughs use AF3 to demonstrate sequence matching, multichain input and MSA settings. Model-specific behavior is described in the guides above.
Saved outputs record their generation date and source versions; fresh runs use the current service and may differ.
Loading examples…
How to interpret your results
If you provide a UniProt ID, inspect its precomputed MSA evidence. If you provide a protein sequence without a reference ID, GreenFold first performs reverse lookup during input preparation to find an eligible reference. The examples below cover both workflows. MSA evidence and reference matching are shared across the supported models; the prepared files and run commands depend on the model.
MSA evidence and previews: P63313
This example retrieves stored alignments directly by UniProt ID P63313. The tables show real canonical source metadata, captured on 29 September 2026, before any sequence changes or model conversion.
| A3M channel | Sequences including query | Depth excluding query | Metadata status |
|---|---|---|---|
| Paired | 4,506 | 4,505 | ok |
| Unpaired | 2,928 | 2,927 | ok |
For the unpaired file, 2,928 records comprise the query plus 2,927 other sequences. Paired and unpaired are separate evidence channels that model adapters consume according to their pairing rules. Row counts describe alignment size; they do not measure Neff, predict accuracy or establish an interaction.
| STO source | Depth including query | Query length | Alignment width | Scan |
|---|---|---|---|---|
| UniRef90 | 3,143 | 44 | 73 | ok |
| MGnify | 140 | 44 | 47 | ok |
| Small BFD | 533 | 44 | 52 | ok |
| UniProt paired search | 4,506 | 44 | 67 | ok |
The 44-residue query occupies an alignment with extra gap columns, so the STO width can exceed 44. STO depth here includes the query, unlike A3M depth. Sources overlap; adding their counts would overestimate unique evidence. ok records a successful metadata scan.
| Record | Aligned sequence |
|---|---|
query | MADKPDMGEIASFDKAKLKKTETQEKNTLPTKETIEQEKRSEIS |
UniRef90_Q5EES1/9-44 | -------AEVKTFDQSNLKHVETEEKNPLPTAATLREELRPEV- |
UniRef90_Q5EES1/122-155 | ----PDVSEVEKFDSSKLKNVTPEVKCHLPTKDVIQEE------ |
In the second record, the first seven - characters mark gaps relative to the query. Uppercase residues occupy aligned columns; lowercase residues, when present in A3M, represent insertions. Open the interactive MSA view and select Preview unpaired A3M to explore more rows and its color legend.
The saved API preview is capped at 2,000 characters and is truncated. The three complete records above illustrate the alignment; the full file contains 2,928 records. Display limits and header filters act on preview records. A STO preview may contain annotations only, so zero parsed rows can still accompany a nonempty full alignment. Download the complete file when needed. Cropping, variation and model conversion can change final files; the tables describe the stored canonical sources.
Sequence-only input: reverse lookup during preparation
Open this interactive result →This input supplies a 44-residue protein sequence without a UniProt ID. GreenFold matches it against indexed references, selects P63313 and infers I10A. The image shows the resulting chain matching report in Prepare, using AF3 as the worked example. The precomputed alignment itself is illustrated in the MSA example above.
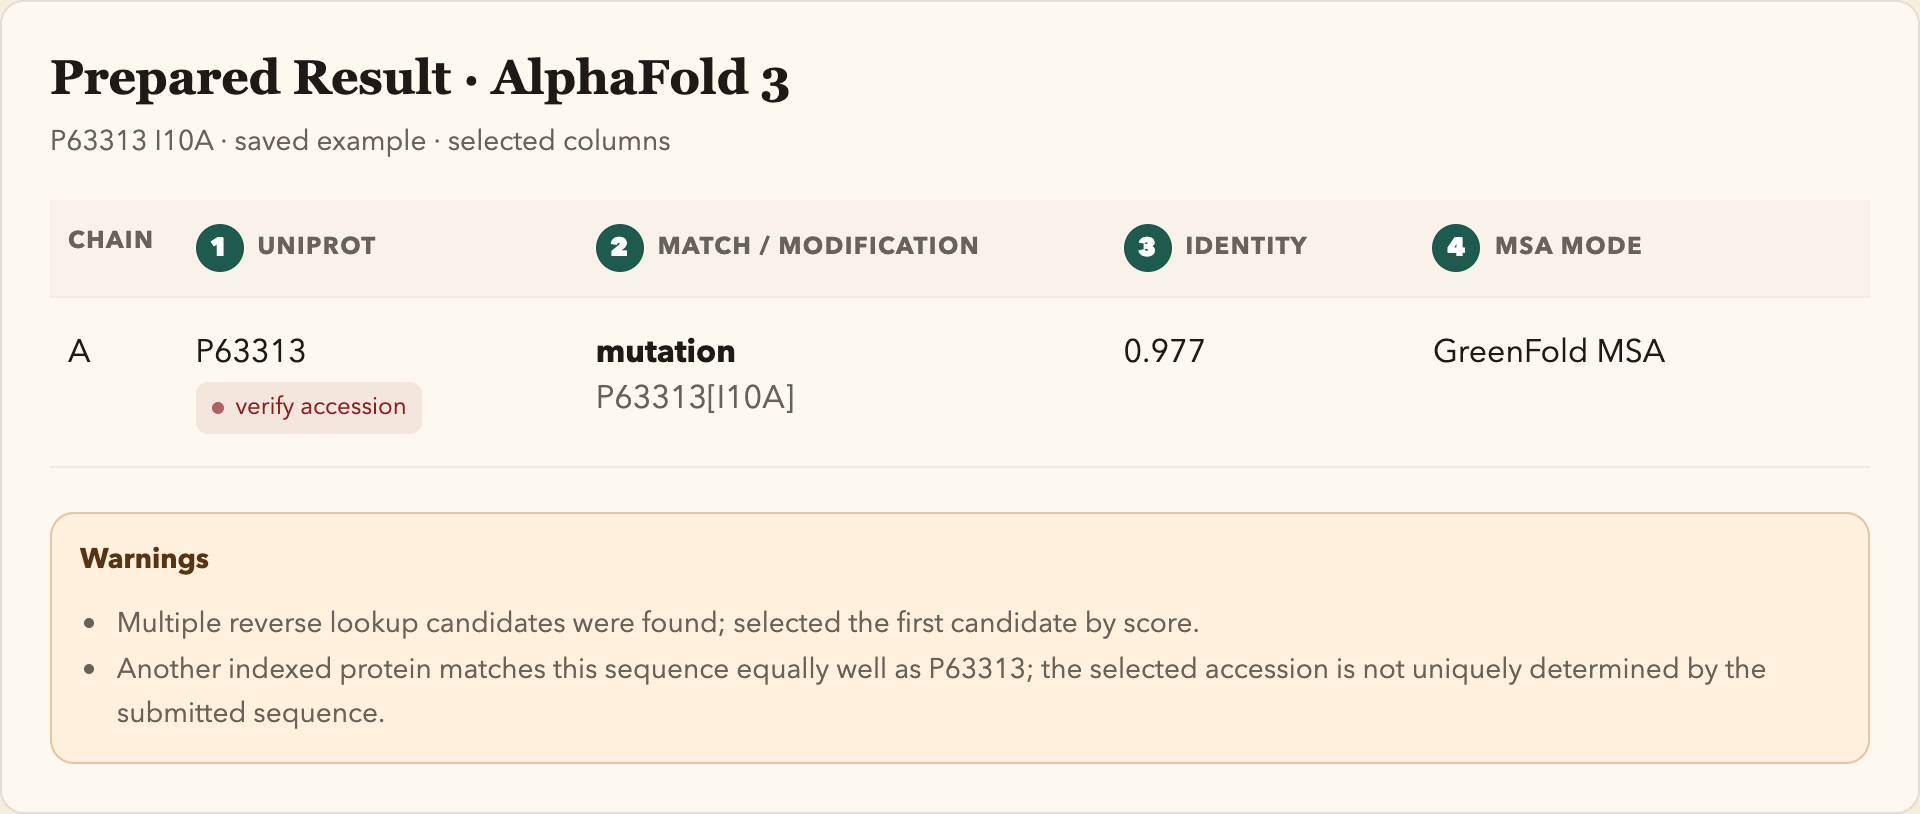
- Reference assignmentP63313 is selected, but verify accession flags an ambiguous match. Check the intended reference.
- Sequence change
I10Ais the inferred substitution at reference position 10. - Identity43/44 ≈ 0.977 measures sequence agreement. A high value can coexist with the warning shown here.
- MSA sourceGreenFold stages MSA evidence from the selected reference with the inferred change applied. Review the assignment before using the package for prediction.
Reference matches and sequence changes
| Result | Meaning and what to check |
|---|---|
| UniProt / exact match | The selected reference for a submitted chain. An exact full-length P63313 match has identity 1.000 and no inferred variation. |
| Identity | Sequence agreement reported by reference matching. In the I10A example, 43 of 44 query residues agree: identity ≈ 0.977. This measures sequence agreement, not structure confidence. |
| Wildtype coverage / cropped MSA | The fraction of the reference span represented by the query. An exact fragment can have identity 1.000 while covering only part of the reference. A cropped MSA keeps the corresponding reference span. |
| Verify accession / ambiguous candidate | The assignment needs review, for example because several entries or paralogs fit the sequence. Check the gene, reference accession and coverage before accepting the selected MSA. |
| Score / candidates | The score ranks sequence alignments; it is not a calibrated probability. Multiple candidates can require manual review even when identity is high. |
| Variation pattern | Changes use the original reference coordinates. I10A replaces residue 10 from I to A; [2-44trunc] retains residues 2–44, with 1-based inclusive coordinates. See the full variation rules for combinations. |
In reverse lookup, inspect all candidates and their warnings. A high identity does not by itself establish that the selected accession is the intended biological target.
What happens to each protein chain
| MSA mode | Meaning and next step |
|---|---|
| GreenFold MSA | The chain uses indexed GreenFold evidence. Check its reference assignment and requested changes. |
| GreenFold-ColabFold partial | Eligible chains reuse GreenFold evidence; missing evidence is completed by the supplied helper on your machine. Run the helper commands before prediction. |
| Native handling / fallback | The chain is left to the model's native workflow. This can require MSA generation in your model environment; the server has not completed that downstream search. |
| Single sequence (query only) | Homologous MSA evidence is explicitly disabled for this chain. Templates are a separate setting; single-sequence mode alone does not disable them. |
| User-provided MSA | Evidence already supplied in the input is retained where supported. Inspect paired and unpaired sources individually in the report when their provenance differs. |
Warnings need action even when a download succeeds. Verify inferred accessions, inspect cropped coverage and follow model-specific fallback/pairing instructions. For mixed multichain inputs, read the fallback guide.
Files in the prepared ZIP
- Prepared native input: the rewritten FASTA, JSON or YAML for your model.
greenfold_input_preparation_report.json: chain assignments, identity, MSA sources, pairing and warnings.bundle_manifest.json: generated and staged files and their source metadata.- MSA files: alignment evidence referenced by the prepared input or runner.
RUN_COMMANDS.mdand runner support: the model-specific next steps. Keep relative paths intact when unpacking.
AlphaFold DB pLDDT describes its displayed reference structure. GreenFold's sequence matching and MSA preparation do not produce a new pLDDT, docking score or predicted complex.
Available web services
This directory reads the current OpenAPI contract and lists every documented operation, including advanced and legacy operations. URLs use the public address https://greenfold.dsdd.one. Replace path placeholders such as {uniprot}, then supply the listed required query/header parameters or body.
Use Swagger to inspect all optional parameters, request schemas and responses, or ReDoc for the full reference. A health response checks lightweight readiness; it does not validate every artifact.
Loading the service directory…
Access services with Python
Download greenfold_api_example.py
This standalone script needs only Python 3.10 or later and requests. It resolves targets, reads metadata, submits a sequence for reverse lookup, checks an A3M query, prepares a native input and parses the ZIP report and manifest. It does not run structure prediction.
python -m pip install requests
python greenfold_api_example.py --uniprot P63313 --model af3 --output-dir greenfold_example
Choose any supported model, or repeat the accession argument for multiple chains:
python greenfold_api_example.py --uniprot P69905 --uniprot P68871 --model boltz
python greenfold_api_example.py --uniprot 'P63313[I10A]' --model af3
python greenfold_api_example.py --uniprot P63313 --sto-job-db mgnify
Look for the canonical accession, MSA record counts, candidate identity and per-chain MSA sources in the console. The output directory contains the downloaded A3M, native input, report, manifest and prepared ZIP. Unpack the ZIP and follow its commands when you are ready to use the model.
To adapt the template, change --uniprot, --model or the explicit native-format block. Use MsaApiClient.get_json, request and download_file with paths from the directory to access other services. POST input preparation accepts raw file contents. Reverse lookup accepts its JSON request schema.
The optional STO example demonstrates submission, status polling with a deadline and download. HTTP 429 retries are bounded. If a job outlives the deadline, use its printed job ID with the status and download services. See the API guide.
Questions & troubleshooting
My protein is not indexed. What should I do?
The GreenFold reference set covers reviewed human proteins. Inspect reverse-lookup candidates and warnings; do not force an unrelated accession. Use the supported ColabFold partial or native workflow for missing evidence. The selected model's guide explains its fallback behavior.
Importing my file reports unsupported fields.
The entity editor regenerates the input and cannot represent every native field. Read the warning and choose Submit the original file instead to preserve the original contents. Editing an entity switches back to regenerated input. Check the preview before submission.
A preview is empty or shows fewer rows than the metadata.
Character limits, display limits and header filters affect previews. Clear the filter, switch to A3M, or download the complete file. The A3M query header can be query rather than the UniProt accession.
Can I disable MSA or template searches?
Single-sequence mode controls homologous MSA use for a protein chain. Skip templates is a separate option with model-specific behavior. Some formats need the supplied runner commands to apply this setting; downloading the native input alone may not suffice. See the model input guide.
How do I handle long downloads or a busy service?
For STO transformations, use the plan service, submit an asynchronous job, poll its status and download when it succeeds. Record the returned job ID. Follow Retry-After when rate limited and report failed jobs rather than treating them as empty results.
What happens after I download a package?
Install or use the chosen model in your own environment, unpack the complete ZIP, inspect warnings and follow RUN_COMMANDS.md. Model code, weights and data have their respective terms. The runner page and runner guide provide further instructions.
Detailed references: AF2 · AF3 · Boltz · OpenFold3 · Protenix.